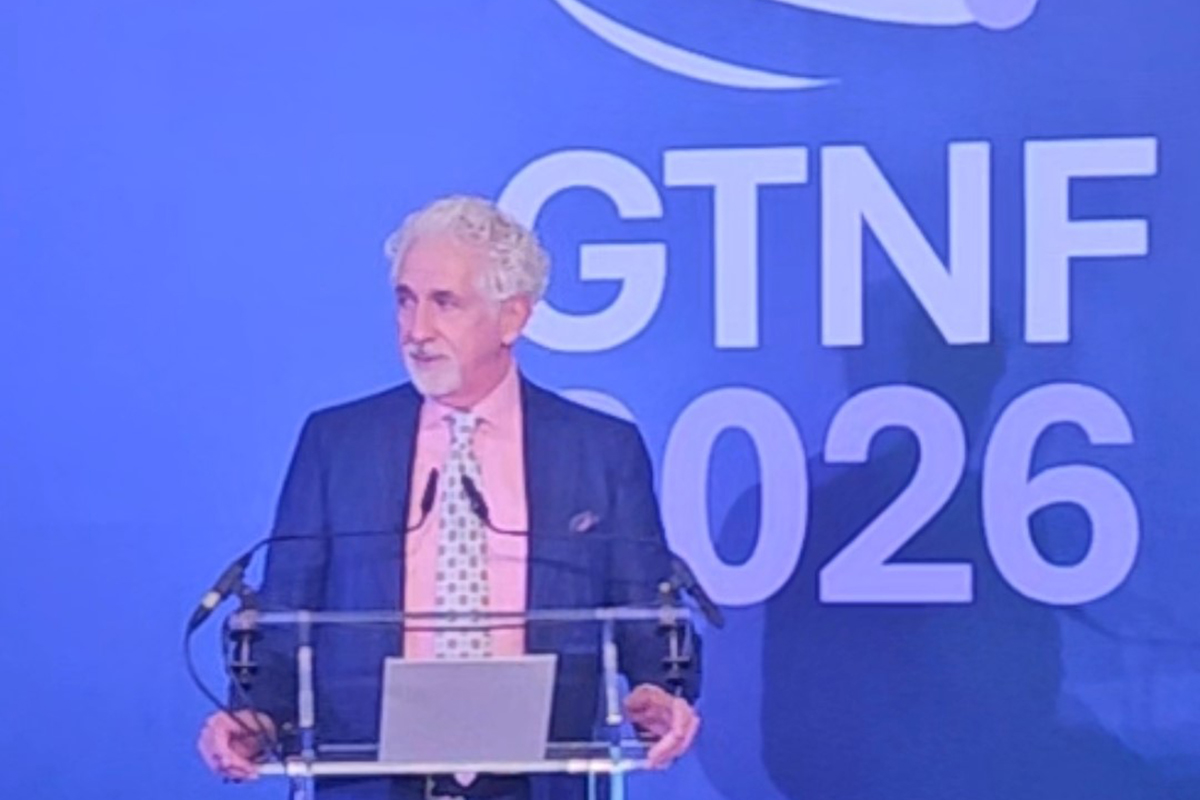
In a keynote address at GTNF 2026 in Lisbon, Dr. Bret Koplow, director of the FDA CTP, said:
I’m here today after having countless talks about the importance of addressing the misperceptions surrounding tobacco harm reduction. And about how we are going to address these issues, including with healthcare providers, so a lot of what we’ve heard here has really resonated.
There’s something particularly fitting about being in Lisbon to discuss the future of tobacco and nicotine. As some of you probably know, centuries ago, Lisbon served as one of the great crossroads of trade, and it was here that Jean Nicot, the French ambassador, obtained tobacco seeds from Portugal in the 16th century and sent them back to Paris, eventually lending his name to the substance that is the very center of our work. Nicotine. Since then, humanity’s understanding of what nicotine and tobacco do to the human body has profoundly changed. With that deepened understanding, we have a responsibility to be good stewards of public health through tobacco product regulation, enforcement, and education, to help people make informed choices about their health and to protect the youth of our nations.
Beth (Oliva) made reference to my background, and I draw on it often in my current role. I’ve sat across from young people whose choices around substances had already begun to shape difficult trajectories that were hard for them to shift. I understand in very direct ways how powerful early exposure to nicotine can be and why the stakes of getting regulation right are not just abstract. I want to spend a few minutes providing perspective about the distinct regulatory context under which we operate in the U.S.
The United States has a dedicated statute governing tobacco product regulation. It’s the Family Smoking Prevention and Tobacco Control Act, which was passed in 2009. It was only 17 years ago that this Act established the Center for Tobacco Products, defined our authority, and created the framework in which we operate. Additionally, it’s only been since 2016 that we’ve had the authority to regulate e-cigarettes and nicotine pouches. Because we’re steeped in this daily, it may seem like we’ve been at this for a long time, but this framework is actually quite new. To a significant extent, differences in tobacco regulation between the U.S. and some other countries is at least in part attributable to the statutory framework that both authorizes and binds our regulatory work. At the same time, the statute gives us a Congressionally defined public health standard against which every new product must be evaluated. This is neither a commercial standard nor simply a safety threshold, but a population-level public health determination. Every authorization we issue requires data that shows the marketing of the product is appropriate for the protection of public health. This standard takes into account a tobacco product’s risks and benefits. It considers people who use higher-risk category tobacco products like cigarettes and their likelihood of obtaining health benefits from switching to this new product. It also considers the risks to vulnerable groups, particularly youth. The standard shapes everything we do in regulating new tobacco products like e-cigarettes and nicotine pouch products, and it’s one of the most consequential features of the U.S. regulatory framework.
The FDA Center for Tobacco Products has a mandate. We regulate cigarettes, smokeless tobacco, cigars, e-cigarettes, nicotine pouches, heated tobacco products, snus, and other products that fall under our definition of a tobacco product, which generally includes any product that contains nicotine from any source that is intended for human consumption, as well as its parts, components, and accessories. Our responsibilities include pre-market compliance, enforcement, public education, rulemaking, and research. But within this broad mandate, I want to be direct about where I believe we can have the most meaningful impact on public health.
There are three things we are currently prioritizing. 1. Preventing youth initiation, especially with combusted products, which are the most harmful and often establish lifelong addiction. Almost 90% of adult daily smokers started smoking by the age of 18, and about 1,500 youth under the age of 18 every day smoke their first cigarette. 2. Helping people in the United States who still smoke cigarettes stop, or switch to products with significantly lower health risks.3. Ensuring people in a position to make those choices, both consumers and the clinicians who advise them, have accurate information about the relative risks of different categories of tobacco products. These three objectives are interconnected. And getting all three right simultaneously is harder than it might seem.
The premarket pathway for new tobacco products has only been operational for a handful of years. The first e-cigarette products were authorized as appropriate for public health less than five years ago. So, in some important respects, we are still finding our footing. In 2018 and 2019, we faced a crisis. We saw an extraordinary, rapid spike in e-cigarette use in young people. Youth use of e-cigarettes doubled in the span of two years, reaching as high as 10.5% of middle school students and 27.5 % of high school students in 2019. It was alarming, unprecedented, and it has left a mark. Around the same time, we also investigated an outbreak of EVALI, and e-cigarette or vaping use associated with lung injury. Even though that was later discovered to be primarily associated with Vitamin E acetate, an additive of illicit THC vaping products, it drastically changed the public’s perception of e-cigarettes. The public health and medical communities’ response when it came to youth was understandable: sound the alarm, restrict access, push back hard. And many of those responses were appropriate at the time, especially given the unknowns around these products then. But, one of the lasting effects of that period, one we’re still contending with, is a widespread and deeply held misperception about what e-cigarettes are and what risks they pose for adults who smoke. Preventing youth use of tobacco products will always remain a priority; however, we must also consider how we can help adults who smoke to reduce the risks of chronic disease and premature death. Today in the U.S., a significant share of adults who smoke and a significant share of physicians believe e-cigarettes are at least as harmful as combusted cigarettes, or that the evidence is too uncertain to conclude otherwise. I know much of this information isn’t new to this sophisticated audience, but I thought it was important to mention it today and say that CTP now endorses these views.
(Audience applauded)
Survey data show that the percentage of adults who correctly understand that FDA-authorized e-cigarettes are generally less harmful than cigarettes actually declined from about 51% in 2012 to under 17% in 2022. That is a striking reversal and a concerning departure from scientific evidence. This matters enormously.
We can review applications, we can authorize products, we can enforce against illegal products. But if the adults who could benefit most from switching don’t believe there is a meaningful difference, or if their doctors won’t recommend switching to FDA-authorized products, we lose much of the public health value of the regulatory work that is so much a focus of our time and effort. Complete cessation of all tobacco use remains the healthiest outcome. That’s our baseline, and it’s unambiguous. But for many adults who try to quit repeatedly and failed repeatedly, completely switching to a lower-risk, FDA-authorized tobacco product could meaningfully reduce their exposure to the toxic and cancer-causing chemicals that make combusted cigarettes so deadly. It’s not a commercial claim; it’s a scientific one. And it’s the harm-reduction basis on which we often issue marketing authorizations.
This brings me to what I think is one of the most important distinctions in our tobacco regulatory landscape: The difference between FDA-authorized products and unauthorized ones. Just last month, a special communication published in the Journal of the American Medical Association by the harm reduction work group of the Society for Research on Nicotine and Tobacco recommends that U.S. clinicians discuss FDA-authorized e-cigarettes as a cessation tool for adults who smoke. That kind of guidance from a peer-reviewed clinical publication is significant. It reflects a growing recognition in the scientific and medical communities that the distinction between authorized and unauthorized products is real and that it matters for public health. Why does it matter? Because we don’t know what’s in unauthorized products. We don’t know how they’re manufactured, what levels of harmful constituents they deliver, or whether they would provide any meaningful benefit to someone trying to switch from cigarettes. The burden of demonstrating that a product meets our public health standard rests with the applicant. And the vast majority of applications we’ve received have not met that burden. To date, we’ve authorized over 100 tobacco products through the Pre-Market Tobacco Product Application review pathway, including e-cigarettes, nicotine pouches, heated tobacco products, and snus, thus extending across nearly all non-combustible categories. Each of these authorizations was only issued after rigorous, multi-disciplinary, scientific review across a wide range of scientific disciplines. The applications for each demonstrated that adult users of high-risk tobacco products could be expected to benefit by completely switching to the new product and that such benefits are offset any identified risk to users and to the general population as a whole. That’s what it means to be FDA authorized. Users of higher-risk tobacco products, who might benefit from switching to one of these authorized products, may not be able to distinguish them from the unauthorized products on the market, too frequently sitting beside them on the same store shelves. We know that cigarette smokers who try and fail to quit using a nicotine replacement therapy, like a nicotine patch or nicotine gum, often conclude such products are ineffective and forego further attempts. I worry about those who may try switching to an unauthorized tobacco product that lacks a meaningful benefit and similarly will wrongly conclude that the entire category offers them no benefit. The opportunity cost of that failed switch is real. Once someone concludes that the category offers them nothing, they may not try again.
No discussion of the U.S. tobacco landscape is complete without addressing flavors. In the wake of 2019, flavors and e-cigarettes became a deeply contested terrain in our country. And with good reason. We know that non-tobacco flavors can be appealing to youth. We know they played a major role in the 2019 youth use surge. And the FDA remains vigilant about that, but we also hear from adults who smoke that they’re drawn to flavors as well. The FDA considers how much the use of a flavored e-cigarette can promote complete switching among the adults who smoke, provided that information is included in that product’s application. That opportunity for risk-reduction is a potential benefit the FDA weighs in its decision-making. In fact, we’ve authorized the marketing of multiple menthol-flavored e-cigarettes partly based on such a claim. Just a few months ago, we authorized the first e-cigarette products in non-tobacco, non-menthol flavors. These products incorporate device access restriction technology, including ID verification, smartphone pairing, and periodic biometric check-ins. These measures effectively limit use to verified adults, 21 and older. The calculus changes where youth access risk is meaningfully mitigated by robust technology. And the approach that the technology that was used in the product that we authorized, that’s just the first. I expect there to be other approaches, some that may not rely on smartphone apps. I know a lot of older smokers in particular may not—my father had a to his last day, had a flip phone still—so your smartphone app isn’t going to work for everyone. It’ll be fingerprint readers that can be programmed at the point of sale by the retailer for the individual user. There may be a whole bunch of different approaches. I am genuinely, genuinely excited about what this could ultimately mean. When combined with existing marketing and point-of-sale restrictions, device access restrictions can provide a novel and robust layer of protection, restricting use to only adults, 21 and older, and that’s the age limit in the United States. And where these flavors that may pose risks to youth are meaningfully mitigated via effective access restrictions, an expanded range of e-cigarette flavors might be more readily made available to adults who smoke, and who want to completely switch from combusted cigarettes. It’s not a silver bullet. But it is a potentially meaningful new tool. And we certainly want to encourage technology and innovation that may similarly work; I want to stress that this is not the only way a company can demonstrate that a non-tobacco flavored e-cigarette meets the public health standard. It remains the case that a non-tobacco flavored e-cigarette may be authorized where the application demonstrates its benefit in helping adults quit smoking outweighs its potential risks.
We don’t have a flavor ban. I know that a lot of people think that we do. There is an additional hurdle in order to be able to demonstrate that the benefits offset the risk to youth. Flavors by themselves are not a problem. We don’t have a concern about flavors. We have a concern about protecting youth. So, one of the things that I think is potentially so exciting about device access restrictions is it takes that out of the equation. I want to be clear, though, that the potential for these new products will not be realized while the U.S. market remains flooded with unauthorized, flavored e-cigarettes. Unauthorized products, many of them flavored, many of them illegally imported into the United States, sit on shelves next to the small number of rigorously reviewed and authorized products. Consumers can’t often tell the difference. Retailers sometimes don’t bother to verify, and the public health benefit of our authorizations is diluted every day these unauthorized products remain on the market. And I know when somebody goes into, often a gas station or convenience store, and they’re looking for a nicotine pouch, let’s say, or an e-cigarette, there’s nothing on the package that tells them which ones are legal for sale and which ones are not; which have been reviewed by the FDA and which ones have not. And so, the consumers are really at a loss and unable to make informed decisions. And we’re talking about various ideas, about helping make better-informed consumers so that the decisions that they make are better.
This is where I want to be direct with this audience because so many of you are in or adjacent to Industry. The unauthorized market is not just a regulatory compliance problem. It’s also a public health problem, and for companies that invested in doing things legally, it’s a competitive fairness problem. Over the past two fiscal years, the FDA and our partners at U.S. Customs and Border Protection have seized millions of unauthorized e-cigarettes worth hundreds of millions of dollars before they entered the country. We’ve dramatically increased import refusals from roughly 100 in fiscal year 2023 to over 9,000 in fiscal year 2025. And that progress is real. It’s not a solution. But it doesn’t mean that trying to mitigate some of this isn’t useful. We know the scale of what enters the U.S. market through unauthorized channels remains significant. We see on the horizon opportunities to do more, with better data sharing, stronger coordination with other federal law enforcement partners, and potentially new regulatory tools. I said earlier that one of our three core priorities is ensuring that consumers and clinicians have accurate information about relative risk. I want to return to that because it’s an area where we have a lot more work ahead. CTP has a webpage that’s dedicated to tobacco harm-reduction education. And we are actively studying the impact of harm-reduction messaging on intended and unintended audiences. We’re looking carefully at expanding our communication on this topic to key audiences while avoiding inadvertently undermining prevention messages to youth. And we’re monitoring the impact of our recent product authorizations on both consumer behavior and public perception. The recent JAMA guidance, the Journal of the American Medical Association guidance that I mentioned earlier, is really encouraging, and if you haven’t read it, I encourage you to do so. I’ve been sending it to a lot of people. When leading scientific societies begin recommending that clinicians discuss FDA-authorized e-cigarettes as part of a harm-reduction conversation, it signals a shift in how the medical community is beginning to engage on this topic. We want to support that shift carefully with evidence and in ways that keep youth protection front and center.
Let me close with this. I’m aware that this international forum draws participants from around the globe. That’s why I traveled to you even after the first two planes I sat on were cancelled for mechanical reasons. The regulatory approach we take in the United States is only one way to go about it. As someone who came into this role with a lot of background in tobacco law, I have a genuine curiosity about how other parts of the world are navigating these crucial questions. And I find conversations like the ones that are happening here genuinely valuable.
This conference has been so stimulating, and so helpful to me personally. I really haven’t been exposed as much as I really probably should have been to how things are being done outside the United States. And the conference has just been so well planned. The agenda has hit every important issue, and one thing that I especially appreciated was that each of you who spoke backed up what you were saying with data. These were not pol-antics. These were scientific data for them, and that’s so crucial for us at CTP. So, thank you to all of the speakers because this was really just wonderfully helpful to us.
The U.S. framework is certainly distinctive. We have a statutory structure that requires us to weigh the public health implications of every product authorization. That structure has led us to develop and deploy rigorous science to confront hard trade-offs and to build institutional capacity that just didn’t exist at all 15 years ago in the United States. We’re still building it. And we’re doing so in a domestic environment that is shaped by a spike in youth use that left a deep weariness about the entire category of e-cigarettes. Awareness, we’re now working to address with evidence, with compassion for adults who use a higher-risk category of tobacco product, who could benefit from switching. Not with alarm.
Many of you in this room represent companies that have invested in or want to invest in developing products that can genuinely benefit public health. Many of these companies may be playing by the rules, submitting applications with real scientific data. Perhaps your products affected youth access restrictions. These companies are partners in the mission that we share. For us, the mission is not about creating profits, though, of course, the pursuit of profits can help drive Innovation—it’s to reduce the harm tobacco has caused for centuries. We have more tools to do that now than we ever did before. We’re continuing to use them to move our strategic priorities forward to advance public health by reducing tobacco-related harm.
Thank you all for welcoming me here.